
For decades, the chemical industry has operated under a prevailing assumption: in the high-temperature world of Partial Oxidation of Methane (POM), metallic nickel is king. As the primary industrial pathway for producing syngas—a vital precursor for synthetic fuels and essential chemical feedstocks—the POM process has been the subject of intense study. Yet, beneath the surface of this fundamental reaction, a mystery has persisted. Scientists have long observed metallic nickel nanoparticles in post-reaction catalysts, leading to the conventional wisdom that these particles are the true active sites.
However, a groundbreaking study published in Nature Catalysis has shattered this paradigm. By peering into the "black box" of the catalyst surface during active operation, an international research team has revealed that the metallic nickel seen after the fact may be a red herring—a product of cooling and reduction rather than the catalyst that actually performs the work. Instead, the true engine of methane conversion is a fleeting, dynamic atomic structure that emerges only under the intense pressure and heat of the reaction itself.
The Chronology of a Catalytic Mystery
The history of POM research is defined by a persistent struggle to reconcile theoretical models with experimental observations. Scientists have known for years that nickel is highly sensitive to its environment; under the reducing conditions of syngas production, it can shift its oxidation state and rearrange its atomic structure with remarkable speed.
Historically, researchers relied on ex situ analysis—examining the catalyst before and after the reaction. This methodology, while standard, created a significant blind spot. Because the catalyst’s state at room temperature is vastly different from its state at high-temperature operating conditions, the "active species" identified in the laboratory were often just the stable remnants left behind once the heat was removed.
The shift toward in situ characterization—observing the catalyst while it is actively driving a chemical reaction—marked a turning point. In this recent study, a collaborative team led by Professors Tao Zhang, Aiqin Wang, and Xiaoyan Liu from the Dalian Institute of Chemical Physics (DICP) of the Chinese Academy of Sciences (CAS), in conjunction with Prof. Wei Liu (DICP), Prof. Tao Yang (Xi’an Jiaotong University), and Prof. Graham J. Hutchings (Cardiff University), set out to solve the enigma. They hypothesized that the real catalytic action was not occurring on static metallic surfaces, but rather on a surface undergoing constant, subtle reconstruction.
Designing the Catalyst: Doing More with Less
To prove their hypothesis, the team moved away from high-metal-loading catalysts. Traditional industrial catalysts often rely on high concentrations of nickel to compensate for low efficiency. The researchers instead synthesized a Ni/Al₂O₃ catalyst using a precise microemulsion method, achieving a nickel loading of only 0.8 wt%.
The performance of this low-loading catalyst was nothing short of exceptional. Despite containing only a fraction of the nickel typically found in industrial counterparts, the material achieved a 92% conversion rate of methane. Furthermore, the selectivity for carbon monoxide (CO) and hydrogen (H₂) reached 87.0%, with a stable H₂/CO molar ratio of approximately 2.0—the "gold standard" for syngas production.
When compared to an 8.0 wt% Ni/Al₂O₃ catalyst prepared via standard impregnation, the 0.8 wt% catalyst showed comparable performance. Even more telling was the comparison to a similarly low-loading (0.8 wt%) catalyst prepared via traditional impregnation, which failed to catalyze the POM process entirely, instead promoting the undesirable complete combustion of methane into carbon dioxide and water. This disparity underscored that it wasn’t just the amount of nickel that mattered, but the specific, refined way in which the atoms were arranged.
Unveiling the [Ni1O4Ni4] Structural Unit
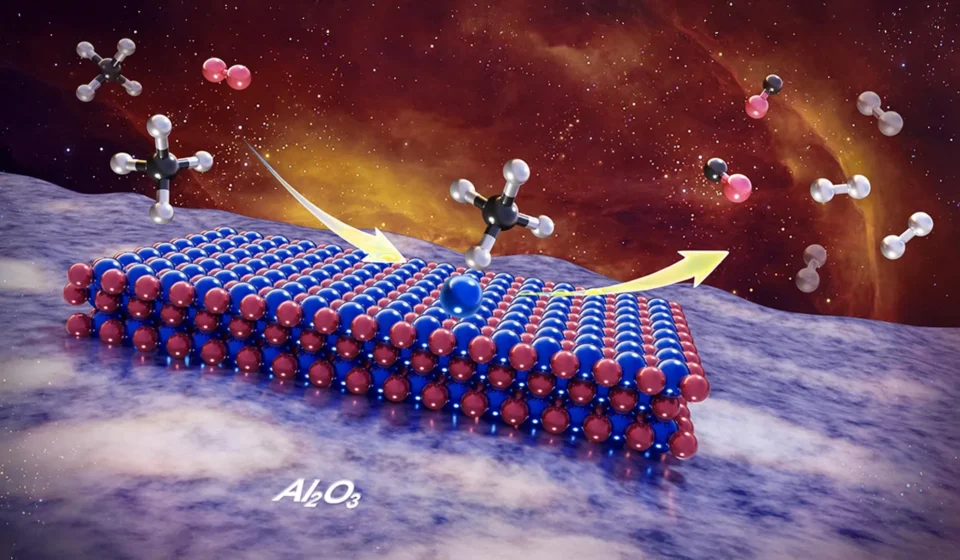
The core of the study lies in the atomic-scale discovery of a specific structural motif. Using advanced in situ techniques, the researchers captured the dynamic formation of a [Ni1O4Ni4] structural unit on the NiO(100) surface. This unit is not a permanent feature of the catalyst’s architecture; rather, it is a transient, reconstructed surface state that only appears under reaction conditions.
To validate that this motif was indeed the active center, the team employed Density Functional Theory (DFT) calculations. The results provided a compelling kinetic explanation for the catalyst’s high performance. The fundamental hurdle in POM is the activation of the methane molecule, specifically the breaking of its robust C-H bonds.
The calculations revealed that the [Ni1O4Ni4] unit acts as a highly efficient catalyst for this step. The activation barrier for C-H bond cleavage on this reconstructed surface was calculated to be a mere 12.5 kcal·mol⁻¹. In contrast, the barrier on an intact NiO(100) surface was a prohibitive 38.5 kcal·mol⁻¹, and even the metallic Ni(111) surface—the long-held candidate for the "best" active site—presented a higher barrier of 15.7 kcal·mol⁻¹.
By lowering this energy requirement, the reconstructed surface allows the reaction to proceed rapidly and selectively. The metallic nickel nanoparticles that researchers usually find after the reaction are, therefore, merely the result of the system returning to a more stable, inactive state once the reaction gases are removed and the temperature drops.
Official Perspectives and Implications for Industry
The implications of these findings are profound for the field of heterogeneous catalysis. For decades, the industry has focused on maximizing metal dispersion or preventing sintering, often assuming that more metal equals more activity. This study suggests that the path to higher efficiency lies in "catalytic engineering"—specifically, designing materials that favor the formation of these dynamic, reconstructed surfaces.
"Our study highlights the critical role of in situ characterization in identifying dynamic active structures under reaction conditions," says Prof. Wei Liu. "Dynamic reconstruction enables low-loading catalysts to achieve high performance, offering new opportunities for the rational design of efficient catalysts while reducing reliance on high metal loadings."
By shifting the focus from the static composition of a catalyst to its dynamic behavior, researchers can begin to design catalysts that "self-optimize" under reaction conditions. This could lead to a new generation of catalysts that are not only more active but also more sustainable, significantly reducing the environmental and economic costs associated with heavy-metal mining and disposal.
A Future of Rational Catalyst Design
The transition from a "trial-and-error" approach to "rational design" is the ultimate goal of modern catalysis. The discovery of the [Ni1O4Ni4] motif provides a template for future research. If scientists can control the surface reconstruction of other transition metal oxides through precise synthesis and structural engineering, the potential for efficiency gains across the chemical industry is immense.
Furthermore, this research serves as a cautionary tale regarding the limitations of standard analytical techniques. As the field moves toward more complex, multi-component systems, the reliance on ex situ observations may prove increasingly insufficient. The ability to "see" the catalyst in its active, transient state is no longer a luxury; it is a necessity for the next wave of chemical innovation.
As the industry faces mounting pressure to decarbonize and increase the efficiency of syngas production, this study offers a clear roadmap. By harnessing the power of dynamic structural reconstruction, we can optimize the processes that sustain our modern world, ensuring that the fuels and chemicals of tomorrow are produced with greater precision, less waste, and a deeper understanding of the atomic dance that makes it all possible.
In the final analysis, the "hidden" structure identified by the DICP team is more than just a scientific curiosity—it is a blueprint for the future of chemical catalysis. By acknowledging that catalysts are not static objects but living, changing participants in a reaction, we open the door to a new era of molecular engineering, where performance is derived not from brute-force chemistry, but from the elegant, atomic-scale choreography of the materials themselves.